by Jonathan Latham PhD and Allison Wilson, PhD
On July 23rd Senator Rand Paul (chair of the Senate’s Committee on Homeland Security and Government affairs, HSGAC) belatedly released the transcript of a Slack messaging group comprising four of the five leading virologists who wrote the now notorious paper ‘The Proximal Origin of SARS-CoV-2’ (Andersen et al., 2020). The four Slack authors are Kristian Andersen (Scripps), Eddie Holmes (University of Sydney), Bob Garry (Tulane University), and Andrew Rambaut (University of Edinburgh). The 1,123 page Slack text, spanning over two years worth of correspondence, can very profitably be read (and has been written up) for several reasons: one being to witness the shameless chicanery and conniving of these four virologists that other scientists, the intelligence community, and politicians chose to rely on, even as the four virologists manipulated them and the scientific peer review system. The transcript can also be read as an unprofessional spectacle of sub-adolescent name calling and profanity. The word F**k is used 180 times; Sh*t is on 241 pages; Prominent scientists, even ones who were agnostic about the lab leak, get referred to by insulting nick-names: ‘Bloomshit’, ‘e-dim’ or ‘E-dumb’, ‘Frankie Bollocks’; while the Paris group of researchers are ‘Parisites’.
But to a virologist, and to anyone determined to solve the COVID-19 origin puzzle itself, it is their secret doubts, the sins against science, against conventional virology, and against logic, of the Slack group–who are experts after all–that reveal what we really needed to know. More than that, as an unexpected bonus, the Slack messages also expose the internal mechanics of their lab leak defense.
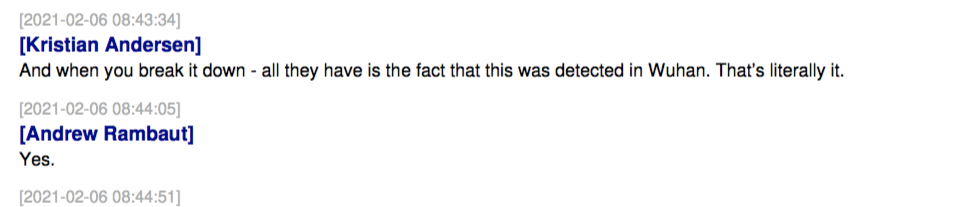
Sure enough, the Slack virologists created a self-righteous bubble, contemptuous rather than respectful of different scientific viewpoints and contradictory evidence. A classic example is this pair of quotes (p491):
And then Bob Garry two months later (p713):
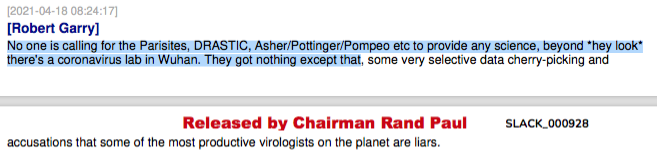
Let us quickly therefore, as a reminder to ourselves, “break it down” for them to illustrate that bubble.
First, an epicentre in Wuhan is several points combined. China has many cities and tens of thousands of live animal markets, but only Wuhan hosts a globally unique coronavirus collecting and research laboratory (the WIV) with a special interest in SARS-related coronaviruses. This laboratory was known even to its own staff for its poor biosecurity (Zhiming, 2019). The city is also nearly 1,000 miles from where the bat coronaviruses closely related to SARS-CoV-2 have been found. Besides the improbability of Wuhan as an epicentre there is the apparently perfect human adaptedness of the SARS-CoV-2 outbreak strain (more on this below). There is also the single phylogenetic origin of the outbreak, a classic hallmarks of lab escapes, and which the Slack authors and others strove hard but unconvincingly to make disappear (Pekar et al., 2022; Bloom, 2023; Cowan, 2025). There are also suspiciously positioned restriction enzyme sites in the SARS-CoV-2 genome (Bruttel et al., (2021), as Kristian Andersen privately noted in early February 2020. Then there are other slightly lesser red flags of a lab origin, such as the inability to find infected intermediate hosts or even susceptible animals being that sold at Wuhan markets.
It is important to highlight the partisan, unbalanced, and frankly anti-scientific thinking pervading the Slack. But the real value in the Slack transcript requires a laser focus on the specific scientific issues that troubled its authors most. And, surprisingly perhaps, the biggest elephant in their bubble was the question of the human adaptedness of the SARS-CoV-2 origin strain.
The Slack authors devoted disproportionate effort to discussing to what extent SARS-CoV-2 was human adapted and comparing this with the adaptation of SARS-CoV-2 strains when inserted in other species; which is another way of determining its adaptedness to humans. The human adaptedness problem became a sticking point for them because it resisted reconciliation with a market origin, and especially it could not be resolved with a market origin in December 2019. For these reasons it eventually became critical to them to dispose of the adaptedness issue. And they did this with an outlandish and unique virological argument, that SARS-CoV-2 was a “generalist virus”. As will be shown, this claim didn’t fit the evidence at all and, furthermore, it would have made SARS-CoV-2 unlike any other coronavirus and arguably any virus.
What thinking lay behind this striking decision?
The troublesome “human adaptedness” question
The word “adapt” appears on 69 Slack pages. Most of these instances refer to viral adaptation, either to humans, or to other hosts by way of comparison.
The underlying explanation for this focus is that the complex lifecycle of viruses requires them to carefully balance multiple competing interests. They typically have to match entry and exit from different cell types with evasion of multiple different forms of host immunity, transmission between hosts, and numerous other parameters that affect their long and short term prospects. Just as we humans must balance our work and home lives, finances, etc., if any significant viral life history factor is removed or altered then the balance between all of them will change. Thus the great fear of virologists who isolate a novel virus is that during amplification in its first petri dish, where the constraints normally imposed by acquired immunity and innate immune cells are absent, the novel virus becomes a non-representative culture-adapted virus, as many older viruses are (Kaul et al., 2007; Shirato et al., 2018). This responsiveness to prevailing conditions comprises a fundamental principle of virology (Simmonds et al., 2019). Because the degree of adaptedness of a virus to its a host is understood to be a highly sensitive indicator of its history, if an unknown virus fails to adapt in a specific host or environment this is as good as a proof that it came from that host and that environment.
Thus Bob Garry wrote in the first Proximal Origin Slack (p45), when the adaptedness question first arose, that evidence of early adaptation or lack of it by SARS-CoV-2 is:
“VERY important. even decisive.”[sic]
In March 2020 the Slack participants plus Ian Lipkin published their Proximal Origins of SARS-CoV-2 in Nature Medicine. It highlighted high infectiousness and high transmissibility as features of the novel virus from Wuhan. It was suggested that this impressive human adaptedness would likely be explained by cryptic circulation. This is the idea that imperfectly adapted precursors of SARS-CoV-2 circulated undetected in humans prior to the detection of known cases, giving them the opportunity to evolve into the better-adapted SARS-CoV-2 that is considered to be the origin strain.
Subsequent research has confirmed the initial adaptedness of SARS-CoV-2 up until the emergence of the Alpha variant, by which date viruses were becoming selected for immune escape (Markov et al., 2023). These assessments used statistical genetics to compare the relative frequencies of synonymous (i.e. silent) mutations (dS) against the frequency of nonsynonymous mutations (dN). The basis of this well-established method is to assume that synonymous mutations are selectively neutral whereas nonsynonymous mutations are more likely to represent adaptations, therefore the relative frequency of the two mutation types provides an estimate of virus maladaptiveness. A poorly adapted virus will generate a high proportion of nonsynonymous mutations as selection pressure forces it to become better adapted. This method (often referred to as dS/dN) is considered uncontroversial and reliable and accordingly multiple research groups concluded that SARS-CoV-2 (in this nine month period) was barely evolving or not evolving at all (Shan et al., 2020; Cagliani et al., 2020; Chaw et al., 2020; Chiara et al., 2021; MacLean et al., 2021; Tai et al., 2022). So whereas according to the zoonosis theory SARS-CoV-2 should be adapted either to the former host (i.e. bats or to an intermediate host) it was, instead, adapted to humans from the moment it was first detected.
Initially, the Slack authors accepted this interpretation. Thus here is Kristian Andersen (p72):

But, in February 2021, Eddie Holmes alerted them to a preprint (later published in Cell) that apparently changed their minds (Kang et al., 2021). Kang et al. suggested (using a novel statistical genetic approach) that the SARS-CoV-2 genome had evolved in humans prior to its first detection and so was not preadapted after all. Simultaneously, (also on pps536-537, which pages are worth reading in full) the Slack authors rejected the accepted dS/dN method. This was for no clear reason and despite having previously made extensive use of it themselves as the first Proximal Origin Slack shows. They reached both of these conclusions by assuming that a novel amino acid in SARS-CoV-2 (A372 in the spike protein, which differs from ancestral bat coronaviruses which feature T at position 372), and first identified by Kang et al., had appeared after the pandemic started. A372 appeared to them proof that the virus had adapted in humans via cryptic circulation.
This reasoning, as Kristian Andersen pointed out to them, wasn’t very solid. It required assuming A372 wasn’t already present in the first SARS-CoV-2 to jump from its previous host (p536):
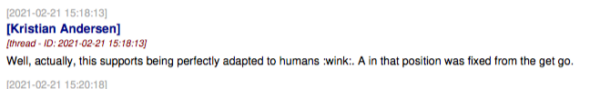
And since genomes with T372 were never found in humans this suggestion of Andersen’s is more consistent with the evidence. But, without offering reasons, the others ignored him.
That wasn’t the end of the Slack preadaptation story. Accepting the Kang result absolutely requires a belief in undetected precirculation. But, in mid 2022 the Slack virologists published two major papers (Pekar et al., 2022 and Worobey et al., 2022) asserting that SARS-CoV-2 originated at the Huanan Seafood Market in December 2019. Exactly when these papers were conceived is not clear from the Slack. But a December 2019 spillover leaves no opportunity for cryptic circulation at all.
To thread this needle, their chosen solution was to reject human adaptedness completely and propose that SARS-CoV-2 was a “generalist virus” with a wide host range, or “pantropic” as Bob Garry called it. The “generalist” virus idea, in contrast to highly human adapted, came from MacLean and colleagues and was enthusiastically adopted (MacLean et al., 2021):
“doubtless SC2 is very pantropic [generalist]” wrote Bob Garry (p602); and a gleeful Eddie Holmes messaged (p715):
Followed by Holmes: “It’s a true generalist” (p781).
And for the remainder of the Slack dialogue, preadaptation to humans (or perfect human adaptation) of the origin strain becomes not just a doubtful proposition but “mind numbingly dumb” (p637), “misinformation” (p695), “nonsense” (p747). In another reversal, their former orthodoxy had become a heresy.
This generalist thesis is important and deserves careful attention. First, by erasing the human adaptedness of the origin strain it removes a formerly immovable obstacle to the market theory. But, second, it is not supportable for two major reasons:
a) The first problem is that the preadaptation idea refers specifically to the origin strain. It is the surprising human adaptedness of the putative outbreak virus that the Slack authors sought to explain. However, the outbreaks they count as evidence for generalism mostly include later strains and novel variants of SARS-CoV-2. Since these later types acquired infectivity towards different mammalian species, such as mice, rats, and white-tailed deer, they are not relevant to the question of whether SARS-CoV-2 emerged as a generalist; in fact they argue against it (Oude Munninck et al., 2021; Shuai et al. 2021; McBride et al., 2023; Schlottau et al., 2020). Similarly, MacLean et al. (originators of the generalist hypothesis) claim infection of pangolins as key evidence of generalism, even though no SARS-CoV-2 infection of a pangolin has ever been reported (MacLean et al., 2021).
In fact, origin or near-origin strains of SARS-CoV-2 infect only a small number of species under experimental settings and transmit between even fewer (Jahid et al., 2024). Those infected outside of labs are almost all mustelids (ferrets/mink or otters) or felines (lions, tigers, panthers, snow leopards, and domestic cats) on farms or in zoos, that is under unnatural and crowded conditions.
b) The second major problem for the generalist hypothesis is that in laboratory experiments with mink and mice, characteristic host-specific mutations immediately appear in the spike and other viral proteins (Leist et al., 2020; Tai et al., 2022; Pulit-Penaloza et al., 2022; Huang et al., 2021). Similarly, outside of labs, e.g. on mink farms and also in wild deer and big cat COVID19 outbreaks, species-specific mutations also appear and for each species are often the same mutations as appear in laboratories. These mutations are adaptations to the new host, thereby demonstrating that in non-human species the initially infecting virus is not perfectly adapted (Tai et al., 2022; Markov et al., 2023; McBride et al., 2023; Iglesias-Caballero, 2024; Bashor et al., 2025).
Thus the contrasting behaviour of SARS-CoV-2 in humans, where adaptations accumulated very slowly, compared to mink, mice, deer, cats, and other hosts where adaptation begins instantaneously, cannot be accounted for by inventing the concept of a generalist virus. Nevertheless, if we continue to consider the medium term, i.e. the first nine months as SARS-CoV-2 spread across the world, a small number of adaptive amino acid mutations did arise. What do they tell us?
What really happened with adaptation?
Fortunately, in 2026, no one need any more depend on statistical genetic estimates to gauge early SARS-CoV-2 adaptation. This is because it is now known which were the key earliest SARS-CoV-2 mutations after emergence in Wuhan and, approximately at least, their functions. And the spoiler is that virologists are hesitant to discuss them in much detail (or it seems fund research on most of them) because none of those identified so far are human-specific adaptations. Rather, they are either transmission mutations or putative transmission mutations. These mutations thus confirm and extend the statistical genetic findings: the SARS-CoV-2 origin strain was indeed fully human adapted and the virologists who dismissed them were wrong to.
In the following analysis we define four mutations as keys to the adaptiveness question. The first was the nucleotide change A23403G. This created the famous Spike protein change referred to as D614G (this notation indicates an amino acid substitution in which G, glycine, replaced D, aspartate, at Spike position 614). the second is C14408U (which gave the substitution nsp12: P323L in which L, leucine, replaced P, proline at position 323 in the nsp12 protein). These first two are non-synonymous changes. In addition to these are the two mutations C241U and C3037U. These are nucleotide changes only and they don’t affect the amino acid sequence of any viral protein (Colson et al., 2024).
These four are critical because they (and only they) share the following characteristics–each arose early in the pandemic (in or before January 2020); each became rapidly fixed (i.e. dominant) in the viral population; and each has been retained through the whole pandemic. For D614G and P323L these unique attributes (earliness, fixation, and retention) can be seen in the figure below, from Flores-Alanis et al. (2024), and they independently corroborate that each mutation strongly boosted viral fitness.
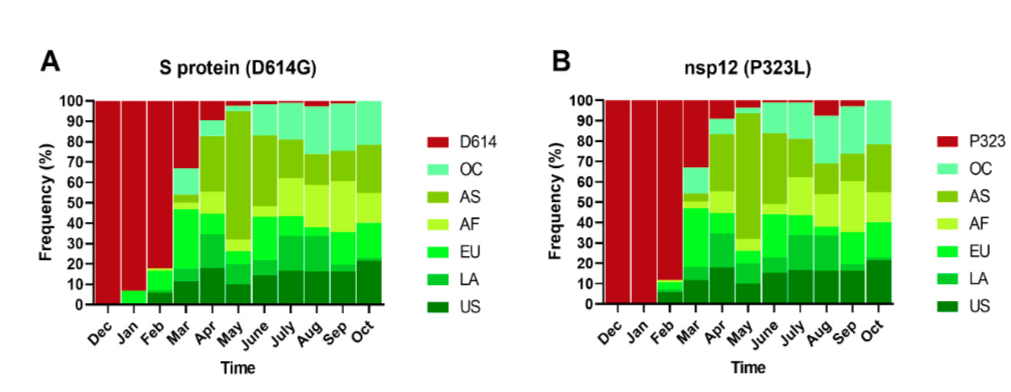
The key question to ask about each is whether they were adaptations specifically to humans or whether they were adaptations to some other selective force operating during the pandemic (with other obvious candidates being epidemic conditions or evasion of the acquired immunity due to prior infection and vaccination building up in the population).
Spike D614G
The most famous of these by far is D614G. This spike mutation has been the subject of hundreds of papers based on early fears that it might evade immunity or be more harmful. These fears went unrealised and, instead, the primary effect of D614G, and why it seems to have been selected, is that viruses with D614G transmit faster and better. Since its effects are always the same, no matter what species is tested, D614G is not a human-specific adaptation (Hou et al., 2020a; Schlottau et al., 2020).
The virological reason why such mutants transmit better was for a while mysterious but is worth knowing since it affirms its role in transmission. D614G has complex effects on the function of the spike protein. These led to the initial expectation it would infect all cells better; but, in keeping with its lack of effect on lung titres, these effects seem to be inconsequential in most tissues, except in the cells of the upper respiratory tract where D614G mutants are more abundant (Hou et al., 2020a; Plante et al., 2021). This advantage explains better transmission and infection; but the mechanistic reason why it grows better only in the upper airways only became clear quite recently. It is that D614G mutants are superior at evading innate immunity, and innate immunity, unsurprisingly, is particularly active in the upper respiratory tract (Mesner et al., 2023). Thus improved resistance to innate immunity leads to better infection in the upper respiratory tract which leads to better person-to-person transmission. This explains why D614 variants were replaced by G614.
P323L
The story of P323L is similar but also different. P323L is a mutation of the viral polymerase. Viruses with L at position 323 replicate faster (but probably with more errors) under standard conditions (Goldswain et al., 2023). The secret to understanding P323L, however, is that it replicates better than P323 at 330C, which is the normal temperature of the nose (Kim et al., 2023). Thus P323L adapts SARS-CoV-2 to the nose, which is the critical organ for transmission and infection. Thus, just like D614G, P323L is a transmission mutation and, just like D614G, it transmits better in all species tested. Thus both P323L and D614G have been tested in hamsters and ferrets and P323L also in macaques (Kim et al., 2023). The bottom line again is that P323L is also not an adaptation specific to humans, but rather an adaptation specific to transmission.
Nose adaptation in an airborne epidemic is critical since most viruses enter a new host by infecting nose cells and equally are exhaled from nose cells; the virus may in fact never reach the lung, or, if the virus does reach the lung, the lung is not the originating site from which transmission occurs (Hou et al., 2020b). Thus, probably contrary to most lay understanding, SARS-CoV-2 goes from nose (person 1) to nose (person 2) to nose (person 3), or, alternatively, from nose to lung to nose (person 1) and to nose to lung to nose (person 2), etc. Whichever pathway is followed, and it may be a mix, adaptation to the tissues of the nasopharynx is a critical requirement for effective transmission. (This leads to an interesting question for the market origin theory; since these mutations are highly beneficial for transmission regardless of species, why they did not arise in the intermediate host?)
C3037U and C241U
Surprisingly perhaps, no credible research has been done on C3037U and only a little is known about C241U. C241U, however, is a mutation of the SL5a hexaloop which is very close to the beginning of the SARS-CoV-2 genome (which is a single stranded RNA). The SL5 region forms a complex structure of folded RNA that is thought to control production of viral proteins and also to control viral packaging. SL5 may act as a switch between these two alternative functions, which are prioritised depending on the stage of infection. Interestingly, C241U partially destabilises the specific loop it is in. By analogy with P323L it is tempting to speculate that this instability enhances functionality at lower temperatures, i.e. in the nose. The bottom line however, is that it is currently not possible to say if C241U or C3037U are human specific adaptations, except to say that they may well not be.
Conclusions
What we learn from analysing how the Slack virologists dealt with the human adaptedness question are two things of critical and enduring importance.
The first is that that the Slack authors, though highly motivated, could find no valid grounds in either logic or virology for dismissing the human adaptedness of the outbreak strain. Rather, their dismissal of human adaptedness was based on pure necessity. The pressure they felt was obscured by the bluster, but the market zoonosis hypothesis rested from the beginning on very slender threads. The strongest evidence linking the outbreak with an animal origin were that (1) multiple December COVID-19 patients had a link to the Huanan Market and (2) some SARS-CoV-2 genomes were found there in surface swab samples. These were facts but their actual import was uncertain. The samples and patients may merely have been representative of Wuhan in December 2019; or they may have resulted entirely from ascertainment bias since China’s CDC initially seemed to assume a market origin. To avoid them being irrelevant the outbreak date had to be pinned to that month. Hence any other finding that threatened a December origin, such as earlier patients or samples, had to be confronted or denied. Obviously, precirculation contradicts a December origin and without precirculation there is no explanation for human adaptedness compatible with a zoonosis. Hence human adaptedness had to go too. Thus the Slack messages reveal at last the strategic rationale underlying the zoonotic defence in the West. This explains the panic (that close followers of the origin story may recall) when on February 25th 2022 the leaders of China’s CDC published a preprint proposing that the Huanan Market in Wuhan was indeed just an ordinary market and SARS-CoV-2 was already widespread by the time it was sampled. (This CCDC manuscript was eventually published as Liu et al., 2023).
The second revelation is the decisiveness of human adaptedness for the origin question. No one, including the Slack virologists, has ever scientifically contradicted the human adaptedness of SARS-CoV-2 as it first appeared in Wuhan. What Kristian Andersen presciently wrote on February 10th, 2020 remains fundamentally accurate today (p24):
“Let’s not forget that what we’ve observed is completely unprecedented as far as I know. Never before has a zoonotic virus jumped into humans and spread through the population like wildfire with this kind of speed. This in itself would require further inquiry.”
“Nothing in biology”, wrote geneticist Theodosius Dobzhansky, “makes sense except in the light of evolution”. And because viruses evolve so rapidly, this axiom applies especially to them: nothing refutes a zoonotic hypothesis like adaptation by SARS-CoV-2 in all animals except humans. With no refutation of it in sight, we are compelled to face as fact that every nucleotide (and not just the spike region) of a genome 30,000 nucleotides in length was unprecedentedly adapted to a supposedly ‘new’ host. And with zero evidence for precirculation in humans one must therefore accept as a fact also that, prior to emergence, SARS-CoV-2 underwent some form of human preadaptation process.
The reasons this hasn’t long been clear are, in part, that the definitive facts have taken time to emerge. But the other part is that senior virologists like the Slack group knew all along that adaptedness could be decisive if the chips didn’t fall their way. And, decisively, they have not — and so the research literature, though rich in evidence, is alternately, coy, silent, or actively misleading about the import of mutations like D614G for the origin question. A classic case is Pulit-Penaloza et al., who infected multiple ferrets with SARS-CoV-2 and deep sequenced the viruses that emerged (2022). Without counting the spike these authors found 18 nonsynonymous mutations that arose multiple times independently and that thereby verify rapid adaptation to the new host; but they failed to note, discuss, or even count them and consigned the entire data set to the supplementary section.
If the scientific case is still not clear the private thoughts and admissions of Bob Garry, Kristian Andersen, Andrew Rambaut, and Eddie Holmes now are. As of today only unreasonable doubt can deny the overwhelming likelihood that a virus leaked from a lab is what the human population of Wuhan was exposed to in 2019. No WIV whistleblower or paper trail, just a close look at the available scientific data, is needed to accept this as overwhelmingly the likeliest cause.
Applying ordinary scientific standards to both the zoonosis hypothesis and the lab leak hypothesis we thus arrive at the real question. It is not whether SARS-CoV-2 was a zoonosis but:
What combination of research, sampling or other processes leads a bat coronavirus to become a fully adapted human respiratory virus?
Here is not the place to propose an answer; but two obvious though very divergent choices are immediately available. There is one that opinion writers in mainstream media or authors with publishing contracts invariably mention when permitted to discuss the origin of SARS-CoV-2; and there is another possibility that they ignore.
References
Andersen, K. G., Rambaut, A., Lipkin, W. I., Holmes, E. C., & Garry, R. F. (2020). The proximal origin of SARS-CoV-2. Nature medicine, 26(4), 450-452.
Bashor, L., Gallichotte, E. N., Galvan, M., Erbeck, K., Croft, L., Stache, K., … & VandeWoude, S. (2025). SARS-CoV-2 within-host population expansion, diversification and adaptation in zoo tigers, lions and hyenas. Nature Communications, 16(1), 11310.
Bloom, J. D. (2023). Association between SARS-CoV-2 and metagenomic content of samples from the Huanan Seafood Market. Virus Evolution, 9(2), vead050.
Bruttel, V., Washburne, A., & VanDongen, A. (2022). Endonuclease fingerprint indicates a synthetic origin of SARS-CoV-2. BioRxiv, 2022-10.
Chiara, M., Horner, D. S., Gissi, C., & Pesole, G. (2021). Comparative genomics reveals early emergence and biased spatiotemporal distribution of SARS-CoV-2. Molecular biology and evolution, 38(6), 2547-2565.
Cagliani, R., Forni, D., Clerici, M., & Sironi, M. (2020). Computational inference of selection underlying the evolution of the novel coronavirus, severe acute respiratory syndrome coronavirus 2. Journal of virology, 94(12), 10-1128.
Chaw, S. M., Tai, J. H., Chen, S. L., Hsieh, C. H., Chang, S. Y., Yeh, S. H., … & Wang, H. Y. (2020). The origin and underlying driving forces of the SARS-CoV-2 outbreak. Journal of biomedical science, 27(1), 73.
Colson, P., Chaudet, H., Delerce, J., Pontarotti, P., Levasseur, A., Fantini, J., … & Raoult, D. (2024). Role of SARS-CoV-2 mutations in the evolution of the COVID-19 pandemic. Journal of Infection, 88(5), 106150.
Ghafari, M., Liu, Q., Dhillon, A., Katzourakis, A., & Weissman, D. B. (2022). Investigating the evolutionary origins of the first three SARS-CoV-2 variants of concern. Frontiers in Virology, 2, 942555.
Goldswain, H., Dong, X., Penrice-Randal, R., Alruwaili, M., Shawli, G. T., Prince, T., … & Hiscox, J. A. (2023). The P323L substitution in the SARS-CoV-2 polymerase (NSP12) confers a selective advantage during infection. Genome biology, 24(1), 47.
Hou, Y. J., Chiba, S., Halfmann, P., Ehre, C., Kuroda, M., Dinnon III, K. H., … & Baric, R. S. (2020a). SARS-CoV-2 D614G variant exhibits efficient replication ex vivo and transmission in vivo. Science, 370(6523), 1464-1468.
Hou, Y. J., Okuda, K., Edwards, C. E., Martinez, D. R., Asakura, T., Dinnon, K. H., … & Baric, R. S. (2020b). SARS-CoV-2 reverse genetics reveals a variable infection gradient in the respiratory tract. cell, 182(2), 429-446.
Huang, K., Zhang, Y., Hui, X., Zhao, Y., Gong, W., Wang, T., … & Jin, M. (2021). Q493K and Q498H substitutions in Spike promote adaptation of SARS-CoV-2 in mice. EBioMedicine, 67.
Iglesias-Caballero, M., Mas, V., Vázquez-Morón, S., Vázquez, M., Camarero-Serrano, S., Cano, O., … & Casas, I. (2024). Genomic context of SARS-CoV-2 outbreaks in farmed mink in spain during pandemic: unveiling host adaptation mechanisms. International Journal of Molecular Sciences, 25(10), 5499.
Jahid, M. J., Bowman, A. S., & Nolting, J. M. (2024). SARS-CoV-2 outbreaks on mink farms—a review of current knowledge on virus infection, spread, spillover, and containment. Viruses, 16(1), 81.
Kang, L., He, G., Sharp, A. K., Wang, X., Brown, A. M., Michalak, P., & Weger-Lucarelli, J. (2021). A selective sweep in the Spike gene has driven SARS-CoV-2 human adaptation. Cell, 184(17), 4392-4400.
Kaul, A., Woerz, I., Meuleman, P., Leroux-Roels, G., & Bartenschlager, R. (2007). Cell culture adaptation of hepatitis C virus and in vivo viability of an adapted variant. Journal of virology, 81(23), 13168-13179.
Kim, S. M., Kim, E. H., Casel, M. A. B., Kim, Y. I., Sun, R., Kwak, M. J., … & Choi, Y. K. (2023). SARS-CoV-2 variants with NSP12 P323L/G671S mutations display enhanced virus replication in ferret upper airways and higher transmissibility. Cell reports, 42(9).
Liu, W. J., Liu, P., Lei, W., Jia, Z., He, X., Shi, W., … & Wu, G. (2024). Surveillance of SARS-CoV-2 at the Huanan seafood market. Nature, 631(8020), 402-408.
MacLean, O. A., Lytras, S., Weaver, S., Singer, J. B., Boni, M. F., Lemey, P., … & Robertson, D. L. (2021). Natural selection in the evolution of SARS-CoV-2 in bats created a generalist virus and highly capable human pathogen. PLoS biology, 19(3), e3001115.
Markov, P. V., Ghafari, M., Beer, M., Lythgoe, K., Simmonds, P., Stilianakis, N. I., & Katzourakis, A. (2023). The evolution of SARS-CoV-2. Nature Reviews Microbiology, 21(6), 361-379.
McBride, D. S., Garushyants, S. K., Franks, J., Magee, A. F., Overend, S. H., Huey, D., … & Bowman, A. S. (2023). Accelerated evolution of SARS-CoV-2 in free-ranging white-tailed deer. Nature Communications, 14(1), 5105.
McCowan, A. (2025) Purported quantitative support for multiple introductions of SARS-CoV-2 into humans is an artefact of an imbalanced hypothesis testing framework. Xarchiv
Mesner, D., Reuschl, A. K., Whelan, M. V., Bronzovich, T., Haider, T., Thorne, L. G., … & Jolly, C. (2023). SARS-CoV-2 evolution influences GBP and IFITM sensitivity. Proceedings of the National Academy of Sciences, 120(5), e2212577120.
Oude Munnink, B. B., Sikkema, R. S., Nieuwenhuijse, D. F., Molenaar, R. J., Munger, E., Molenkamp, R., … & Koopmans, M. P. (2021). Transmission of SARS-CoV-2 on mink farms between humans and mink and back to humans. Science, 371(6525), 172-177.
Pekar, J. E., Magee, A., Parker, E., Moshiri, N., Izhikevich, K., Havens, J. L., … & Wertheim, J. O. (2022). The molecular epidemiology of multiple zoonotic origins of SARS-CoV-2. Science, 377(6609), 960-966.
Plante, J.A., Liu, Y., Liu, J. et al. Spike mutation D614G alters SARS-CoV-2 fitness. Nature 592, 116–121 (2021).
Pulit-Penaloza, Joanna A., Jessica A. Belser, Xiangjie Sun, Claudia Pappas, Nicole Brock, Troy J. Kieran, Jana M. Ritter et al. Comparative assessment of severe acute respiratory syndrome coronavirus 2 variants in the ferret model. MBio 13, no. 5 (2022): e02421-22.
Schlottau, K., Rissmann, M., Graaf, A., Schön, J., Sehl, J., Wylezich, C., … & Beer, M. (2020). SARS-CoV-2 in fruit bats, ferrets, pigs, and chickens: an experimental transmission study. The Lancet Microbe, 1(5), e218-e225.
Shirato, K., Kawase, M., & Matsuyama, S. (2018). Wild-type human coronaviruses prefer cell-surface TMPRSS2 to endosomal cathepsins for cell entry. Virology, 517, 9-15.
Simmonds, P., Aiewsakun, P., & Katzourakis, A. (2019). Prisoners of war—host adaptation and its constraints on virus evolution: OPINION. Nature Reviews Microbiology, 17(5), 321-328.
Tai, J. H., Sun, H. Y., Tseng, Y. C., Li, G., Chang, S. Y., Yeh, S. H., … & Wang, H. Y. (2022). Contrasting patterns in the early stage of SARS-CoV-2 evolution between humans and minks. Molecular Biology and Evolution, 39(9), msac156.
Worobey, M., Levy, J. I., Malpica Serrano, L., Crits-Christoph, A., Pekar, J. E., Goldstein, S. A., … & Andersen, K. G. (2022). The Huanan Seafood Wholesale Market in Wuhan was the early epicenter of the COVID-19 pandemic. Science, 377(6609), 951-959.
Zhan, S. H., Deverman, B. E., & Chan, Y. A. (2020). SARS-CoV-2 is well adapted for humans. What does this mean for re-emergence? bioRxiv. doi: https://doi.org/10.1101/2020.05.01.073262
Zhiming, Y. (2019). Current status and future challenges of high-level biosafety laboratories in China. Journal of Biosafety and Biosecurity, 1(2), 123-127.
Zhou, P., Yang, X. L., Wang, X. G., Hu, B., Zhang, L., Zhang, W., … & Chen, H. D. (2020). A pneumonia outbreak associated with a new coronavirus of probable bat origin. nature, 579(7798), 270-273
If this article was useful to you please consider sharing it with your networks.

Outstanding publication. Leaves me encouraged that a new generation of scientists will continue to discredit the conflicted actions of the ‘old guard’. Accountability is likely a long ways off but sunlight will guide thé way by showing the real conspiracy.
God day. I am NOT a scientist, but do read a lot.
In an article published in the beginning of the plandamic by a South American source: A fishing trawler left with 12 healthy men on board for about 14 days. When they returned, a virus check was done. One sailor positive!
How can that be?????
What spread throughout the world was an e-mail to the laboratories informing the to use the PCR test in this way to identify SARS-CoV-2! The person who wrote the recipe, based on Kary Mullis’ work was Prof. Christian Drosten, backed by Prof. Lothar Wieler from the Robert Koch Institute.
I rest my case.
This is new to us. Thank you. Jonathan
The main paper claiming to disprove a “single phylogenetic origin” was Pekar et al. 2022 in Science, as you cited. Its conclusion was based on a simple error in the Bayesian logic they purported to use. The error can be minimally fixed using only the data, model, and simulations of the paper itself. Repairing the error reverses the conclusion. See
https://econjwatch.org/articles/an-article-in-science-on-covid-origins-contains-a-fundamental-error
Dear Michael,
We meant to cite your paper but lost track of it while travelling this week. Thank you.
.. these unique attributes (earliness, fixation, and retention) can be seen in the figure below, from Flores-Alanis et al. (2024),
-> The figure is missing
Also remember the online searchable slack at https://nemonominem.github.io/PO_Slack/, which combines the 2 releases.
Thank you Gilles. The fig was indeed missing. And I didnt even know about the searchable Slack.
Great work. How relevant is Jim Haslam’s observation that the original SARS2 strain only transmitted in five New World (lab) animals but not in any Old World or Asian animals? Haslam thinks this suggests US virus development. Others said superior human adaptation nullifies this argument. But in a lab leak scenario, how is such extreme human adaptation even explained?
Haslam:
“Through this process, we discovered the WA1 strain transmits efficiently in just five species on this planet: American mink, American deer, American deer mice, American lab bats (i.e., Egyptian fruit bats), Syrian hamsters”
https://jimhaslam.substack.com/p/an-open-letter-to-fauci
I wasnt aware of it and cannot understand this Haslam position. Egyptian fruit bats are from Africa, Syrian Hamsters are from the middle east and he forgot ferrets (which are effectively identical to mink and have the same ACE2 protein) and are from europe, and he forgot various felines (from Europe, Asia, and Africa). So there is a fundamental problem with his observation, is there not? It is true that these species transmit with variable efficiency but that is difficult to estimate/compare in a lab situation (compared to zoo settings or with domestic cats), especially since the experimenters in these cases used very initial high infectious doses. But also I dont really see how or why a virus would be tailored to “asian” or “American species” or whatever species. It should be considered that quite a few commentators on the origin question, including the “there is no virus crowd”, and quite prominent “critics”, may be just spamming you.
Thanks Jonathan. Jim’s observation was that Chinese (lab) bats apparently weren’t infected or didn’t transmit the virus:
“Why only North American (lab) animals (not Chinese bats), can become both infected and ‘efficiently’ transmit SARS2?” https://jimhaslam.substack.com/p/2-why-did-the-batman-fly-into-wuhan
If this is a lab virus and it was perfectly adapted to human transmission, is this because of human cell culture experiments/passaging? Is this consistent with the Darpa Defuse proposal?
We consider the lab leak to close to a certainty. The suspicious restriction sites imply engineering and they are quite compelling. However, the DEFUSE grant does not specifically address human adaptedness except by specifying a “human-adapted FCS”. The problem is that there is no such thing as a specifically human FCS, or an FCS specific to any species. So that statement is hubris, like other parts of the DARPA grant and genetic engineering research in general it operates in vacuum where viruses rarely and sometimes never get put back in a realistic host to test whether they really are more pathogenic (or whatever). This flaw enables specious claims and the DARPA grant is pretty bold in what it claims to be able to do but it isnt clear to us these goals are truly feasible.
Further, some human adaptedness is presumably possible through cell culture/passaging of a bat virus but whether this would create a viable pathogen in humans is not known. But quite likely not for reasons set out in the article.
So, partly for these reasons, we think it is a lab leak but mostly not a lab made virus and the DARPA grant is a red herring.
It’s not necessary that the DEFUSE FCS be unique to humans but it would be some human FCS familiar to the researchers. The RRAR of ENaC fits the bill.
Together with the RE site pattern, independent of fitness constraints, closely fitting DEFUSE plans that makes some engineering likely.
Haslam’s arguments about where good wild hosts live make no sense. The wild virus could not have come from North America so potential wild hosts here are irrelevant. Maybe his argument about lab Syrian hamsters is relevant, but it doesn’t seem very compelling.
The RBD looks just like a pangolin virus. Fauci’s diary says that CIA intercepts showed a Beijing lab sending pangolin coronaviruses to Wuhan for use in experiments with transgenic mice.
Dear Michael
Thanks to you (and Sam) for the constructive comments.
I don’t fully agree the RBD “looks just like a pangolin virus”.
The closest RBD (and the closest spike), according to alignments is BANAL-52. RaTG13 is equally close if you ignore the short section which RaTG13 lost due to recombination after the split with SARS-CoV-2 (Boni et al., 2020; https://www.nature.com/articles/s41564-020-0771-4). RaTG13, BANAL-52, and SARS-CoV-2 are phylogenetically equidistant. So none is the exact progenitor of SARS-CoV-2. For these reasons the known pangolin covs are red herrings too. Now, closer pangolin Covs might exist but there seems no reason ((beyond pangolins being potential intermediate hosts) to pull pangolin covs into the conversation when there are these closer bat covs. Small disclaimer: there are newer bat covs (from Hassanin) that are pretty close that I havent integrated into this scheme but if anything they would make pangolin covs even less relevant.
Thanks Jonathan. You guys have done outstanding work, also on Ebola if I remember correctly.
1) Concerning Defuse, Alex Washburne (who did the restriction site analysis) still thinks it’s very relevant:
“The 2018 DEFUSE grant proposed to modify a bat sarbecovirus in Wuhan by adding a furin cleavage site (FCS) in a reverse genetics system of such a virus. SARS-CoV-2 emerged in Wuhan with a furin cleavage site and a pattern of cutting/pasting sites in its genome, sites modified by a hotspot of silent mutations bioengineers like to use, consistent with a reverse genetics system made by enzymes used in a 2017 paper by several authors of the aforementioned grant. FOIA’d drafts of this grant also contained more specific plans – designating the specific location for the FCS insertion between the S1 and S2 subunits of the S gene (the same place the FCS is found) and including order forms for one of the aforementioned enzymes hypothesized to have been used to synthesize the reverse genetics system that caused a pandemic.” https://alexwasburne.substack.com/p/everyone-is-wrong-about-covid-origins
This does sound quit damning, doesn’t it?
2) Concerning Ratg13 and Banal52, Flo Debarre argues the Banal52/Laos discovery made the Mojiang mine irrelevant. Do you agree or is this a premature conclusion? I remember Ratg13 was your own starting point, but your theory probably doesn’t depend on it?
“sampled in Laos, the BANALs include the now closest known relative of SARS-CoV-2. In 2020 and in the first half of 2021, however, the closest known relative was RaTG13, sampled in the Mojiang mine in Yunnan. The Mojiang mine was therefore big in leaker stories – and still is for those who ignore new data.” https://pandemonium.hypotheses.org/2372
3) Concerning Haslam, I guess he assumes WIV would have tested the virus on Chinese horseshoe bats, but apparently those aren’t SARS2 hosts, while US lab animals are. But maybe WIV would have done something else with the virus?
4) The fact that SARS2 wasn’t found in the wild, combined with the fact that the virus looks very much engineered (both FCS and restriction sites), makes it look like this virus was assembled from scratch, not just collected in the wild and passaged through some cell cultures?
Thanks for these questions Sam
Some of them are answered in the article.
Point 1:These points are largely redundant with each other and the FCS has a function that is location-dependent, if it was anywhere else it would kill the virus. We accept that the restriction sites indicate manipulation but they are independent of engineering in other ways. A mine virus may have been reverse engineered to put restriction sites in it and this is how covs are standardly treated, even ones isolated from the wild. It doesnt tell one whether other parts were engineered or not, but it is evidence of a lab leak. Re the FCS, we think there are a lot of reasons to think it evolved and wasnt engineered. a) it is only a partially functional FCS. Why would an engineeer add a partially functional FCS? Makes no sense. b) if it was added for a science experiment no scientist has ever or would ever add it as an insertion, and nor with a proline. Therefore it probably wasnt added for those reasons. Perhaps it was added for other reasons? It is hard to rule out of course bioweaponeering, but its not a very successful weapon.c) We showed in a poster that there is a parsimonious and plausible evolutionary route.
Point 2: I followed your link to Flo’s blog but I couldnt find that exact argument or anything similar. You know that something isnt true when Flo argues the point. But seriously, BANAL-52 emphasises the importance of the mine because it shows that bona fide plausible ancestors of SARS2 are out there and found quite near to the mine (given that it was sampled 8 yrs later) as are all the other ancestors of SARS-CoV-2. The mine is close to the centre of origin of SARS-C0V-2 viruses is all you need to know. Flo’s text you quote doesnt specify an actual argument and I originally investigated the Slack to see whether the Slack virologists believed that RaTG13 discredited our miner hypothesis because they also believeed (correctly) it cannot be a direct ancestor of SARS-CoV-2. It turns out that Holmes stated this several times but Bob Garry also pointed out in the slack text that its hardly likely that sampling in the mine would detect the exact virus the miners got. So they did appreciate that this argument, which I dissected in detail here was bogus after all.
Point4: Our position is that it was collected from the miners and never passaged at all. Plus if you want to argue it was purely engineered you also need to explain how spike mutations S50L,A372T H519N were acquired. I dont think that can be done.
Thanks Jonathan.
Concerning the suboptimal FCS and strange proline, this has been countered by some. These are quotes from Deigin, VanDongen, Alina, and some others:
“Considerations for engineered origin: 1. CGG is human optimized 2. Polybasic for 10x more cutting effic 3. Proline in PRRA like in MERS: PRSVR 4. CG(GCGGG)yields a FauI site useful for RFLP screening of FCS kicking out 5. Which also explains alanine (GCA, as TCA was split)”
“I think the Proline was there by design, because it was part of a peptide sequence intended to be an immune adjuvant. PRR is certainly popular in patents. ”
“if you introduce a BsaXI site at the position it is, with the goal of using it to insert an FCS, it creates the Proline. It’s forced. Not a design choice. It also explains the Proline! Because the BsaXI site needs to end in CTCC. The reading frame is dictated by the TNS amino acid sequence, resulting in CCx for the next amino acid, always encoding a Proline. So the Proline was not the results of (un)intelligent design, as hotly debated earlier, but a consequence of the Golden gate, ‘no see-um’, cloning strategy. Except that the ‘invisibility’ failed and backfired, because @breakfast_dogs still ‘saw-um’ That is right: if BsaXI was used to create this insert, it would force a Proline at position 681.”
“Well, possibly because you have a bioinformatic prediction that inserting a P with your RRA will create O-linked glycans on the serines that flank your newly created FCS? Also, the PAA in RmYN02/BANAL strains could have inspired the PRRA insertion.”
“This preprint is relevant for some questions that NO proponents have raised: *Why a proline? *Why RRAR? *How would they know to add O-linked glycosylation sites (Thr678 and Ser686)? A: Because MERS.”
“The MERS hypothesis would explain why they used a “non-canonical” FCS, and what the proline was doing there. They were trying to simulate a SARS-like virus getting an FCS like the one in MERS.”
“The only relevant restriction site in SARS2 is BsaXI. BsaXI flanks the out-of-frame furin cleavage site (PRRAR) and forces the Proline, It was published by Baric at UNC in 2017 and BsaXI was designed to be “retained”. Baric was asked by US military of any restriction sites, but said no”
“It also explains the Proline! Because the BsaXI site needs to end in CTCC. The reading frame is dictated by the TNS amino acid sequence, resulting in CCx for the next amino acid, always encoding a Proline”
“proline and out-of-frame insertion pointing to BsaXI GoldenGate cloning site”
“In the final paper from Zhengli Shi prior to the pandemic, a collaboration with PLA members from the Academy of Military Medical Sciences, experiment with the non-canonical, proline-containing FCS of MERS.”
Hi Sam (and Michael) and thank you for putting together this useful list.
It seems to us there are two separate questions for an engineering hypothesis (as opposed to a passaging hypothesis) to answer with regard to the inserted PRRA: 1) why would anyone add a furin site, and specifically this furin site? and 2) why would anyone also add proline (P681), which lies outside the core furin consensus domain?
As a context for this discussion we would like to point out that the specific FCS which SARS-CoV-2 contains is suboptimal in respect of cleavage efficiency (all studies demonstrate partial cleavage of the spike in culture); but it is not suboptimal for the virus itself. Proving this, attempts to “optimize” the site by increasing its cleavage efficiency do increase cleavage but they reduce entry (in cell culture). This is consistent also with the conservation of the PRRA in the first nine months, in which time there was ample opportunity to select for increased cleavage but it didnt happen. During the epidemic, only with the advent of novel variants did the P681 residue become superfluous (lost from the population) and even then its replacement by H or R did not result in fully cleaved spikes, though H/R681 mutants are slightly better cleaved than P681.
Given all this it is interesting to speculate what incomplete cleavage may contribute to the entry process of wild-type SARS-CoV-2 particles. It is known that cleavage is required for early pathway cell entry and this ability strongly promotes entry into lung cells, but cleavage also destabilises spike trimers which become prone to premature shedding (becoming incompetent for entry thereby). Thus it might be expected that incompletely cleaved trimers should be unable to enter via the early pathway, yet the PRRAR FCS promotes early pathway entry while giving only 60-70% cleavage efficiency. Our suggestion for resolving this paradox, consistent with spike trimer structure, is that cleavage of one spike protomer at S1/S2 results in cleavage of its neighbour at S2′ (and not of itself) by TMPRSS2; and for the third and final cleavage the whole structure is loose enough that lack of precleavage at the third S1/S2 site is moot. Hence not all S1/S2 sites in any one trimer need to be cleaved for early entry. Thus the 60-70% cleavage observed in vivo for wild-type SARS-CoV-2 equates to most spike trimers having two cleaved protomers and one uncleaved protomer. In sum, incomplete cleavage of spike trimers represents a sweet spot, it is useful in stabilising spike trimers but it still promotes early pathway entry.
Here then are our answers to Sam and Michaels questions:
Looking at these suggestions they boil down to five categories of theory for the P and the furin site:
1) BsaXI forces it:
2) P creates O-glycan
3) P creates an antigenic site
4) Because MERS
5) Because ENaC
Answer to 1) Assuming the desire to add a furin site and a restriction enzyme cleavage site, BsaXI insertion does require a proline but the introducing of restriction sites by researchers is pretty much invariantly done so as not to change the amino acid sequence (which might disrupt the virus), so it’s possible but not very likely that the ‘need’ for a restriction site introduction”forced” anything. This proposal also underestimates the significance of the proline in year 1 of the pandemic. Prior to the appearance of the alpha strain (in Oct 2020) the P681 residue was very highly conserved. This conservation of the proline (P681) indicates that the virus positively selected for the maintenance of P681. If a researcher had put it there and then the virus was indifferent or found it lowered fitness it would have been lost or poorly conserved. Since this didn’t happen it strongly suggests that P681 was positively adaptive and not research driven.
Answer to 2) Why would a researcher introduce an O-glycan into a live virus? “Because MERS” seems hardly a reason. Assuming it is accepted that the P of MERS creates O-glycans it isn’t clear why these would be useful in SARS-CoV-2. And the second problem is identical to the point discussed above: why should the virus would maintain a site introduced for scientific or similar reasons? And thirdly, this rationale offers no explanation for the FCS itself.
Answer to 3) Why would a researcher introduce an antigenic site into a live virus? This logic implies the creators of SARS-CoV-2 were making a vaccine. The problem begged here is: why introduce an artificial (and therefore non antiviral) antigenic site in order to trigger antibodies against a bona fide virus? And the second problem is: what kind of vaccine project necessitates a respiratory virus (which SARS-CoV-2 is) rather than a bat virus? Neither bat vaccination nor human vaccination requires that, and if SARS-CoV-2 is the product of engineering it would require a large investment of time and money to have made it. This would have included mutating T372 (to A), N519 (to H), and other changes to give respiratory adaptation, plus, separately, human adaptation. Hence making SARS-CoV-2 was a massive work project for no reason.
Answer to 4) First, the inserted PRRA residues are different from the PRSV found in MERS; “distinctly MERS-like” (or “because MERS”) is therefore a weak rationale. This proposition, to make sense, needs a separate explanation for the two amino acid differences (PRSV versus PRRA). I.e. why should the introduction of the residues PRSV eventuate in SARS-CoV-2 with a PRRA insertion? We cannot conceive of a reason. Second, and related to this, PRSVR is a very weak FCS (10% cleavage rate), much weaker than PRRAR (60-70%), vs 100% for a consensus RXK/RR. It is unclear why anyone would add a weak FCS given that all previously inserted FCS’s are strong ones (RXK/RR) or similar.
Answer to 5) The first problem with arguing that PRRA derives from the human gene ENaC-alpha is that ENaC-alpha has no Proline. Thus, as with answer 4, it is necessary to propose a separate reason for P681. As already mentioned restriction sites do not constitute a strong reason, thus we consider the proline to constitute a fundamental objection.
In summary, we think that the lack of precedents for an inserted PRRA and its proven adaptedness during the pandemic constitute powerful reasons against an engineered explanation for the FCS of SARS-CoV-2. Added to this, insertion (rather than a substitution as in other scientifically added furin sites) and the lack of a clear rationale for ever adding a furin site (as was pointed out in the margin of the DEFUSE grant itself, an FCS added to SARS-CoV itself never enhanced cell entry) between them make an engineered PRRA pretty implausible (even though it is technically very feasible).
Maybe the question for you is: How would we know the difference between
1) an evolved (found) virus that was given restriction sites to facilitate manipulation and
2) one that either was a bat virus engineered to become human and lung adapted for an unknown purpose or
3) was a bat virus engineered roughly according to the DEFUSE grant?
And what we have attempted to draw attention to here is that, whereas the DEFUSE grant somewhat matches SARS-CoV-2, it also exhibits major differences.
Concerning Flo, the article is about the 7896 clade and the fact that it wasn’t covered up (by WIV), but her point is that since the discovery of Banal, Ratg13 and the Mojiang mine are no longer that relevant anyway (as they were in 2020). Obviously she supports a natural origin, which I agree is very unlikely, but I think she has debunked many incorrect claims made by Drastic and some others, mainly concerning WIV.
There is a simple answer to this question. The Mojiang mine is relevant because it is associated with an outbreak, not specifically because RaTG13 was found there. Finding RaTG13 there helps understand the mine as in the region where SARS-CoV-2 emerged. So Flo is aiming at a straw man.
She is right they are somewhat less relevant but 1) the location of all similar viruses is relevant to working out the location of the spillover, however conceived. Second, if you chooose to consider the miner evolution hypothesis that we favour then the mine is rather relevant. It is fine for Flo to reason away the miner theory (if she can) but it isnt reasonable just to pretend it doesnt exist, which is what she and a lot of other people inexplicably do (exhibits include Alina Chan in her book, Steven Quay in his book, RFK Jr in his book, everyone else in their books, except Elaine Dewar in her book, who puts it front and centre). It’s worth asking how and why Alina C, Steven Quay, Andrew Rambaut, Eddie Holmes, Flo Debarre, can all be on the same illogical and wrong page. What is so radioactive about evoln in the miners? Is DEFUSE a limited hangout? Etc.
“Plus if you want to argue it was purely engineered you also need to explain how spike mutations S50L,A372T H519N were acquired. I dont think that can be done.”
Would be interesting if you could take this up with / challenge those who propose an engineered virus, if they have an answer to this or not.
Jonathan- I’m puzzled about your remarks on why the PRRA insert wouldn’t have been engineered. As others have said, it’s distinctly MERS-like, and MERS was being studied in the same lab. MERS also has extra length there, like the insertion. DEFUSE did not say that a maximally efficient FCS would be tried but only a human-specific one. The RRAR is straight out of ENaC. It’s also known that maximally efficient is not the same as maximally pathogenic, as in FIPV.
You mention that an FCS in other locations would be unfit. I was wondering roughly how much position freedom the fitness constraint allows. 0 nt? 3 nt?
Hi Michael
Sorry for the slow reply.
As it happens we have done some detailed work on R residues in bat Covs and in sarbecoviruses and merbecoviruses there is a quite small window which is determined mainly by the 3-D structure of the spike protein. Two rules are evident:
1) more widely the cleavage site has on the one hand to be positioned so that cleavage enhances RBD exposure before binding to ACE2 and on the other so as to promote cleavage at S2′ by TMPRSS2 or other surface proteases after binding.
2) More narrowly the R has to be exposed on the surface. A buried R wont cleave no matter what and the Rs of sarbecoviruses are all positioned on exposed loops (whether in bats or not). See this diagram, which isolates the R of BANAL-52 and shows it in relation to the buried region (the lower half of the diagram) and the upper half which is the surface exposed loop. Like many of them the exposed loop in the vicinity of the R is unstructured, hence the dashed lines. The structure shown is extracted from a representative Protein Data Base entry:
https://www.independentsciencenews.org/wp-content/uploads/2026/09/8XYH-knot-structure_03-scaled.png
Well, Flo’s main point in that article is that the 7896 clade (“the other 8”) wasn’t “covered up” by WIV (as suggested by Drastic, Alina and some others), and that seems to be correct. The Mojiang mine is just a secondary point related to the fact that Ratg13 no longer has the unique importance it had in 2020, when it was the closest relative. (When it comes to Banal/Laos, there is also the fact that the US military was sampling these viruses…) Flo has a terribly caustic style and I certainly don’t agree with her natural origin thesis, but I do believe she has debunked many incorrect anti-WIV lab leak claims.
Michael, yes the MERS and ENAC connection are really quite troubling and may well suggest research/engineering.
Concerning the FCS, my understanding is that it doesn’t have to be at the S1/S2 boundary but can be elsewhere in the spike protein, especially within S2. SARS2 has an S1/S2 cleavage site, but other protease cleavage sites can affect activation of the spike and membrane fusion.
In fact Flo Debarre argued that Defuse didn’t plan to insert an S1/S2 FCS, but Defuse drafts showed otherwise. Flo also argued that the “superfluous” proline speaks against engineering, but as shown by Deigin/Chan/VanDongen, it actually supports engineering.
Sam- I’be been trying to check how tight the evolutionary constraints are on the position of the near-S1/S2 FCS. The tertiary structure looks like there might be a few amino acid leeway but so far indels in that vicinity are extremely rare. So I don’t think the precision of the position match to DEFUSE plans provides much extra evidence for a lab leak.
As you say, there are other non-FCS routes for such viruses (e.g. SARS) so the general match to DEFUSE plans tends to support LL but not strongly due to the clear fitness advantages of the FCS.
Jonathan- Thanks for that reply. I scrounged around a little and found a structure picture like that and figured maybe there was a 1 or 2 AA range that would work. It seems that there are almost no indels found nearby so maybe even that overestimates the leeway. This means that the FCS hitting exactly the DEFUSE spot provides very little extra evidence of engineering.
100% agree. When Coronavirus researchers report introducing an FCS they invariably modify the upstream few residues by turning them into R or K. Unfortunately we may not know about failures.
This is a substantial piece that deserves careful scrutiny. Let me offer a critical review of both the article and the comments.
## Critical Assessment of the Article
**Strengths:**
1. **Valuable documentary evidence**: The Slack transcript is a genuine primary source, and examining how virologists privately reasoned versus published is methodologically sound. The contrast between early recognition of “adaptation” as “VERY important. even decisive” (Garry, p45) and later dismissal is genuinely striking.
2. **Careful attention to what counts as evidence**: The article distinguishes between “human-specific” adaptations and “transmission” adaptations—a conceptually important move. The analysis of D614G and P323L as transmission rather than human-specific mutations is plausible.
3. **Specificity on key mutations**: Rather than hand-waving about “adaptation,” the article identifies concrete mutations and asks what they actually demonstrate.
**Significant Weaknesses:**
1. **Overconfidence on the adaptation question**: The article claims human adaptedness is “decisive” against zoonosis, but this conflates several distinct claims:
– A virus CAN be preadapted to humans through intermediate species
– A virus CAN evolve through early cryptic circulation
– A virus CAN jump into humans while carrying adaptations to features humans share with other mammals (respiratory tract structure, ACE2 receptor presence in bats)
The article addresses some of these but not with enough nuance. The “unprecedented” speed Andersen noted could reflect many things besides lab origin—including founder effects, transmission bottlenecks, or the particular ecological conditions of Wuhan.
2. **The “generalist virus” critique is somewhat strawman-ish**: The article sets up the generalist hypothesis as obviously wrong, yet:
– Some viruses genuinely do have broad host ranges (e.g., certain bunyaviruses)
– The article’s evidence against generalism (lab animals acquiring species-specific mutations) doesn’t necessarily refute that the *origin* strain was generalist
– The critique relies heavily on what later variants acquired, not what the original strain was
3. **Assumes strategic motivation too readily**: The article interprets the Slack virologists’ reasoning shift as deliberate deception in service of a preferred hypothesis. But scientists often:
– Change views when encountering new data (Kang et al., 2021)
– Rationalize away concerns that create cognitive dissonance
– Make errors in reasoning without conscious dishonesty
The profanity and name-calling is unprofessional, but doesn’t prove the science is wrong.
4. **The mutation analysis has gaps**:
– D614G and P323L being transmission mutations doesn’t prove the origin strain wasn’t already human-adapted
– “No human-specific adaptations identified so far” ≠ “no human-specific adaptations existed”
– The article acknowledges C3037U and C241U are poorly characterized, weakening the strength of conclusions
5. **Selective citation of phylogenetic work**: The article cites Pekar et al. (2022) as defending market origin, but the real phylogenetic question—where SARS-CoV-2 *actually* came from—remains genuinely uncertain. The article doesn’t adequately engage with competing phylogenetic interpretations.
## Critical Assessment of the Comments
**Michael Weissman’s pushback:**
– Makes a substantive technical point about Bayesian logic errors in Pekar et al. (worth investigating)
– Presses Latham on why the FCS proline would be retained if engineered (fair question)
– Latham’s response is detailed but somewhat circular—arguing the proline’s conservation proves natural evolution, which assumes what needs to be proven
**The Engineering Hypothesis Discussion:**
The lengthy exchange about furin cleavage sites, BsaXI, ENaC, and MERS reveals genuine technical uncertainty:
– The DEFUSE match is suggestive but not conclusive
– The proline issue genuinely cuts both ways (could support engineering via cloning constraints OR natural evolution via conservation)
– Latham’s point that engineered FCS insertions elsewhere “are not precedented” is an argument from absence
**Jonathan Latham’s Responses:**
– Generally substantive and engaging with technical details
– But occasionally shifts ground (e.g., on whether the Mojiang mine matters)
– His “mysterious” emphasis on why others ignore the miner hypothesis while he favors it suggests this is a central commitment—worth interrogating whether this biases his interpretation
## The Bigger Picture
The article makes a **real and important point**: the Slack messages do show prominent virologists privately struggled with human adaptedness and then changed their tune. This is worth taking seriously for what it reveals about scientific reasoning under pressure.
**However**, the article overstates its case by claiming this settles the origin question. What it actually shows is:
– Lab leak is plausible (possibly more so than zoonosis)
– Natural zoonosis is harder to explain than defenders admitted
– The scientific consensus formation was influenced by professional/institutional factors
But it doesn’t prove lab leak. The adaptedness argument is suggestive, not decisive. And the furin cleavage site evidence (restriction sites, DEFUSE match) remains ambiguous—consistent with engineering but not conclusive.
The most intellectually honest position after reading this would be: **”Lab leak is now the less-dismissed hypothesis, but we still don’t know the true origin with confidence.”**
Claudeai- Very cool response.On the “generalist” question, however, I think you misstated the logic. Multiple spillovers from humans to about 8 other species all show fast early amino acid evolution, as did SARS 1in humans. That contrasts with SARS 2. It is a strong indication of probable human adaptation before the earliest official cases. Whether that would be in a lab or remote rural area is another question to be decided by other evidence.
Good day. Thank you for this article.
Rambault and Drosten both spoke to a belief about a long pre-detection period in Wuhan. In other words, the entity named SARS-CoV-2 had been circulating for a period prior to the launch of a test.
What is your explanation for why this circulation was not “showing up” in mortality or morbidity data? (Or perhaps you believe it was?)
Hi Jessica
Sorry not to post your question immediately.
I am not sure which of their comments you are referring to but most commentators agree on about July to Oct as the likeliest emergence date of SARS-CoV-2 based on phylogentics and available genomes. Xia et al., 2021 is arguably the best of these papers, since its based on the most genomes. It gives a date of mid august 2019. However, such a date doesnt necessarily imply a lot of cases and so mortality in Wuhan and elsewhere might not be noticeable. On the other hand there may have been plenty of cases and there is evidence eg for cases in Italy and elsewhere well before Dec 2019 (cited in Xia). Germane to this article is whether such cases/circulation could have allowed preadaptation. It is hard to answer this question with any certainty but its worth considering the possibile permutations. The likeliest one to my mind is that adaptation to humans likely required quite a few mutations since humans and bats are very distant relatives. And if ferrets are any judge there were at least 20 mutations required to adapt SARS-CoV-2 to them (Pulit-Penaloza et al, 2023). One imagines that, at a minimum, a similar scale of mutations is required for either bat to human adaptation or even putative raccoon dog to human adaptation. That is a lot of mutations. So while in a putative period of adaptation in humans (eg going back to august 2019 or before) it is plausible to suppose that a handful of mutation steps occurred but evaded detection I think it is very unlikely indeed that dozens of mutations appeared independently in Wuhan and yet only one adapted strain was ultimately detected. Against this it seems that the authorities in the West and in China made every possible effort to not find viruses prior to Dec 2019. We wrote about this here: Delete, Deny, and Destroy: Chinese and Western Strategies To Erase COVID’s Origin Are Being Exposed By Independent Research though I would suggest we didnt then understand the strategic value to them of these efforts.